High-throughput SNP genotyping relies on well-controlled, reproducible PCR steps that deliver unambiguous allele discrimination at scale. PACE® (PCR Allelic Competitive Extension) genotyping chemistry, developed by 3CR Bioscience, is a homogeneous, endpoint PCR technology designed specifically for robust SNP and Indel detection using fluorescence-based data capture.
This article provides a step-by-step technical walkthrough of the PCR steps involved in a standard PACE genotyping reaction, from DNA preparation through thermal cycling and endpoint fluorescence detection. The workflow is comparable to KASP™ and TaqMan™, but with distinct advantages in assay simplicity, cost efficiency, and throughput.
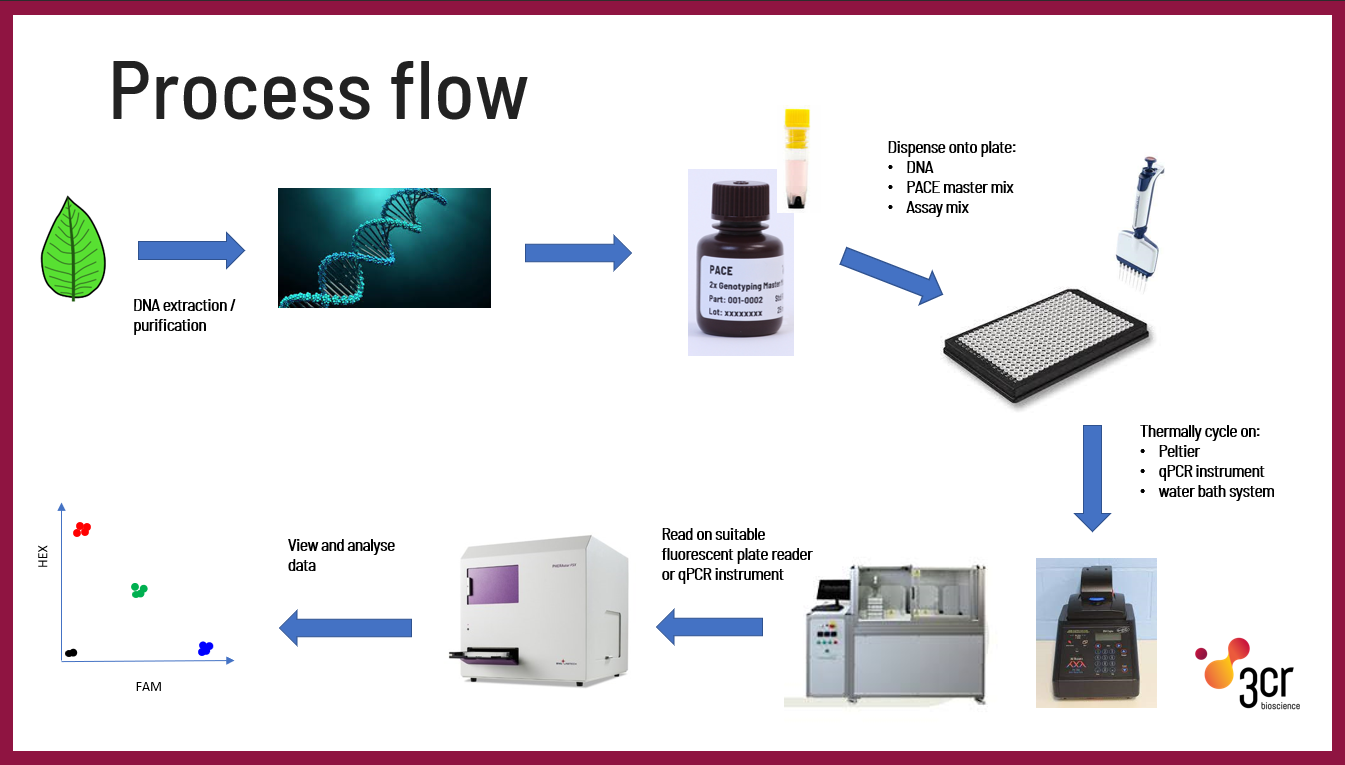
Every PCR reaction, whatever the application, is built on the same three steps: denaturation, annealing and extension. This article explains those three steps first, with the temperatures, times and cycle numbers that apply in practice, and then follows the complete eight-step workflow of a PACE genotyping reaction from DNA preparation through to endpoint fluorescence detection.
The Three Steps of PCR
A PCR reaction repeats three temperature steps in a cycle. Each cycle doubles the number of copies of the target sequence, so 30 cycles produce in the order of a billion copies from a single template molecule.
- Step 1, denaturation. The reaction is heated to 94 to 98 °C for 20 to 30 seconds. The hydrogen bonds between the two DNA strands break and the double helix separates into two single strands.
- Step 2, annealing. The temperature is lowered to 50 to 65 °C for 20 to 60 seconds. Primers bind to their complementary sequences on the single strands. The annealing temperature is the parameter that governs specificity: too low and primers bind at unintended sites, too high and they do not bind at all.
- Step 3, extension. The temperature is raised to 72 °C, or held at the annealing temperature in two-step protocols. DNA polymerase extends each primer along the template, synthesising the complementary strand. Taq polymerase adds roughly 1,000 bases per minute.
A reaction also begins with a single enzyme activation step and usually ends with a final extension. The table below gives the parameters that apply to a standard reaction and to a PACE genotyping reaction.
| PCR step | PACE genotyping | Purpose |
| Enzyme activation | 94 °C, 15 min, 1 cycle | Activates the hot start polymerase |
| Denaturation | 94 °C, 20 s | Separates the double strand |
| Annealing | 65 to 57 °C touchdown, then 57 °C, 60 s | Primers bind, determines specificity |
| Extension | combined with annealing | Polymerase synthesises the new strand |
| Cycles | 10 touchdown plus 30 | Each cycle doubles the product |
| Final step | endpoint read at 40 °C or below | Completes products, or reads the result |
The PACE protocol differs from a standard reaction in two respects. It uses a touchdown phase, in which the annealing temperature is reduced by 0.8 °C per cycle over the first ten cycles, and it combines annealing and extension in a single step. Both serve the same purpose, namely clean separation between the two alleles.
DNA input and purity: what a PCR reaction needs
A PCR reaction fails more often because of the template than because of the cycling parameters. As a guide, 1 to 10 ng of genomic DNA per reaction is sufficient, with larger genomes requiring more. An A260/A280 ratio of 1.8 or above indicates DNA that is free of protein, and an A260/A230 ratio of 2.0 or above indicates DNA that is free of salts and polysaccharides. Plant samples in particular carry polyphenols and polysaccharides that inhibit the polymerase. The extraction method therefore determines whether the three PCR steps can work at all.
Overview of PACE® Genotyping Chemistry
PACE genotyping is an allele-specific, PCR-based technology that combines competitive primer extension with a universal fluorescent reporting cassette. Unlike probe-based chemistries, PACE assays require only unlabelled primers, reducing assay cost and simplifying design.

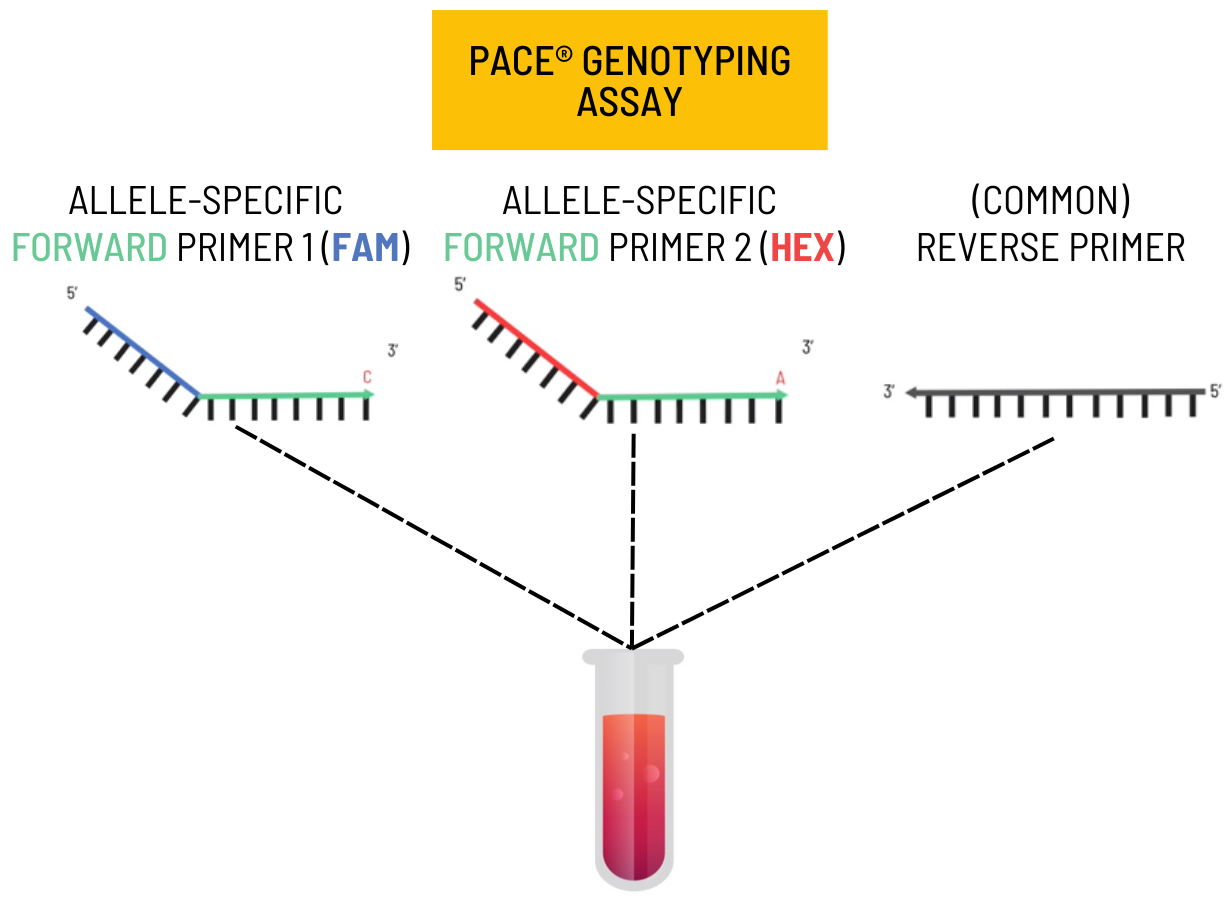
A complete PACE Genotyping Reaction consists of:
PACE Genotyping Assay
- Two allele-specific forward primers
- One common reverse primer
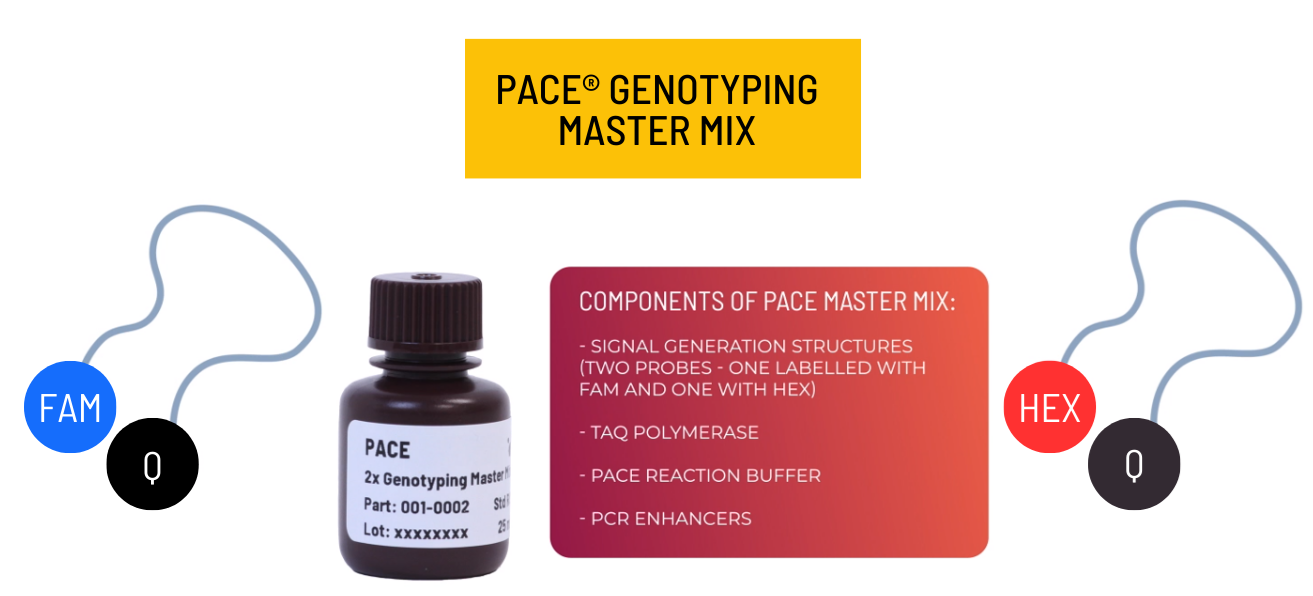
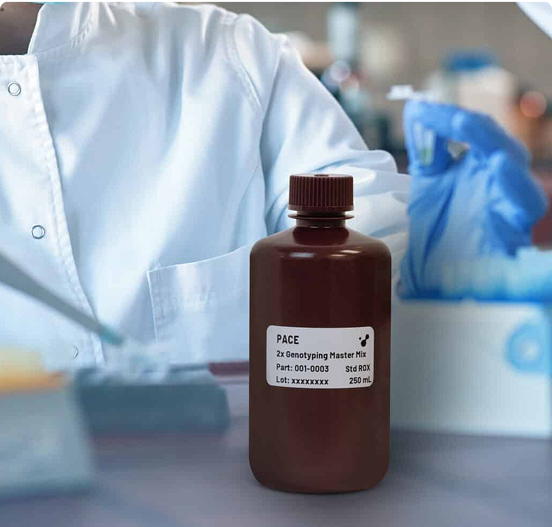
PACE Genotyping Master Mix (2×)
- Engineered Taq DNA polymerase
- Universal quenched fluorescent reporting cassette (FAM and HEX)
- dNTPs, buffer, performance enhancers
- MgCl₂ (4.4 mM at 2×; 2.2 mM at 1×)
- Passive reference dye (ROX; multiple formulations available)
When combined with template DNA, these components create a closed-tube, endpoint PCR assay ideally suited for high-throughput workflows.
PCR Step 1: Template DNA Quality and Preparation
The first of the critical PCR steps is ensuring suitable DNA quality and quantity.
DNA Input Recommendations
- 1–10 ng genomic DNA per reaction well
- Required input varies with genome size (larger genomes require more DNA)
PACE chemistry is particularly tolerant of crudely extracted DNA, which makes it highly suitable for large-scale genotyping projects where purification is impractical.
Key Considerations
- Test a DNA dilution series to identify optimal concentration
- Crude lysates may contain PCR inhibitors
- If inhibition occurs:
- Dilute samples to reduce inhibitor concentration
- Consider PACE® 2.0 Genotyping Master Mix
- Final EDTA concentration must not exceed 0.1 mM
PCR Step 2: Experimental Controls for Genotype Confidence
Incorporating appropriate controls is an essential PCR step for validating data quality.
Recommended Controls
No-Template Controls (NTCs)
- Contain hydration buffer only
- Should cluster near the origin of fluorescence plots
Positive Controls (if available)
- DNA samples of known genotype
- Should cluster in expected allele positions
The absence of amplification in NTCs confirms that fluorescence signals observed in test samples arise from true allele-specific PCR rather than contamination or non-specific amplification.
PCR Step 3: Arraying Template DNA
Template DNA may be arrayed in either hydrated or dry format, depending on throughput requirements.
Hydrated DNA
- Suitable for low-sample numbers
- Faster setup
- More susceptible to evaporation across plate edges
Dried DNA (Recommended for High Throughput)
- Improves plate uniformity and clustering
- Eliminates evaporation-induced concentration variability
- Enables ultra-low reaction volumes (e.g. 1 µL total volume)
Dried DNA is stable long-term at ambient temperature and improves reproducibility in large-scale workflows.
PCR Step 4: PACE Genotyping Reaction Assembly
Reaction assembly is a key PCR step that directly affects assay performance.
PACE Genotyping Master Mix must always be used at a final 1× concentration.
Example Reaction Setup (384-Well Plate, 5 µL Total Volume)
| Component | Volume |
| 2× PACE Genotyping Master Mix | 2.5 µL |
| PACE Genotyping Assay | 0.069 µL |
| Template DNA | 2.5 µL |
| Total Volume | 5.0 µL |
PACE master mixes are compatible with all reaction volumes and plate formats.
PCR Step 5: Reaction Dispensing and Plate Sealing
Once assembled, the PACE Genotyping Reaction is dispensed into the PCR plate.
Best practice includes:
- Using automation for large sample numbers
- Sealing plates with an optically clear seal
- Centrifuging plates to ensure all components are located at the bottom of the wells
PCR Step 6: Thermal Cycling and Allelic Competitive Extension
Thermal cycling is the core of the PACE PCR workflow, enabling allele-specific primer binding and signal generation.
Standard PACE Thermal Cycling Protocol
Step 1: Enzyme Activation
- 94 °C for 15 minutes
- 1 cycle
Step 2: Touchdown PCR (Optimise specificity)
- 94 °C for 20 seconds
- 65–57 °C for 60 seconds
- Decrease annealing temperature by 0.8 °C per cycle
- 10 cycles
Step 3: Amplification
- 94 °C for 20 seconds
- 57 °C for 60 seconds
- 30 cycles
The touchdown phase improves allele discrimination and reduces non-specific amplification—particularly beneficial for challenging SNPs or Indels.

PCR Step 7: Fluorescent Signal Generation and Detection
PACE genotyping uses a universal fluorescent reporting cassette, removing the need for allele-specific probes.
Mechanism of Signal Generation
- Allele-specific primers bind at the SNP position
- Primer tail sequences are incorporated into the amplicon
- Complementary tail sequences are generated during PCR
- Quenched fluorescent cassettes bind these complements
- Fluorescence (FAM or HEX) is released upon binding
Genotype Interpretation
- Homozygous → single fluorescence signal
- Heterozygous → mixed fluorescence signal

PCR Step 8: Endpoint Fluorescence Reading
After thermal cycling, fluorescence is measured in endpoint mode using:
- Fluorescent plate readers, or
- qPCR instruments capable of FAM and HEX detection
Key Detection Conditions
- Read fluorescence at ≤40 °C
- Use a temperature-controlled read step when using qPCR instruments
If genotype clusters are not well defined:
- Perform 3 additional PCR cycles
- Re-read fluorescence
- Repeat up to a maximum of four times
This flexibility ensures tight, well-separated genotype clusters.
PCR Steps at Scale: High-Throughput and Multiplexing
PACE chemistry is engineered for high-throughput PCR workflows, offering consistent performance across large sample sets.
For even greater efficiency, PACE® Multiplex Master Mix enables:
- Detection of up to four targets in a single reaction
- Seamless combination of existing assays
- Reduced DNA input, consumables, and labour
No optimisation is required, making multiplexing accessible even in large genotyping pipelines.
Assay Design Support and Optimisation
3CR Bioscience offers a free PACE® assay design service, providing:
- Allele-specific primer design
- Expert optimisation guidance
- Optional in-house validation
Assays are delivered as primer sequences, allowing customers to source oligos from their preferred manufacturer.
Frequently Asked Questions About PCR Steps
What are the three steps of PCR?
Denaturation at 94 to 98 °C separates the double strand, annealing at 50 to 65 °C lets the primers bind, and extension at 72 °C lets the polymerase synthesise the new strand. The three steps repeat for 25 to 35 cycles.
How long does one PCR cycle take?
Between 1.5 and 3 minutes, depending on the protocol and the length of the target sequence. A full run of 35 cycles plus activation and final extension takes roughly 1.5 to 2.5 hours.
How many PCR cycles are needed?
25 to 35 cycles for a standard reaction. A PACE genotyping reaction uses 10 touchdown cycles followed by 30 amplification cycles.
How much DNA is needed for PCR?
1 to 10 ng of genomic DNA per reaction. Larger genomes require more. The final EDTA concentration must stay below 0.1 mM.
What temperature is used for annealing?
Typically 50 to 65 °C, set 3 to 5 °C below the melting temperature of the primers. In a touchdown protocol the temperature starts high and falls by 0.8 °C per cycle.
Why does PCR fail?
Most often because of the template, not the cycling programme. Inhibitors from crude lysates, too little DNA and contamination are the three most common causes.
Conclusion: Reliable PCR Steps for Scalable Genotyping
From DNA preparation through endpoint fluorescence detection, the PCR steps in PACE genotyping are designed for accuracy, scalability, and ease of use. By combining allele-specific competitive PCR with a universal fluorescent reporting system, PACE delivers robust SNP and Indel genotyping without the complexity or cost of probe-based assays.
Whether supporting breeding programmes, population genetics, or large-scale marker validation, PACE provides a streamlined PCR workflow built for modern, high-throughput genotyping demands.